
Cystic Fibrosis: 7 Essential Facts You Should Know
Cystic fibrosis explained: the CFTR gene, inheritance, symptoms, newborn screening and sweat test, CFTR modulators, life expectancy and daily care.
Introduction

Cystic fibrosis (CF) is passed down through families, and it makes the body produce mucus that is far thicker and stickier than normal, clogging the airways, the gut and other organs. It affects about 40,000 people in the United States and an estimated 105,000 diagnosed people across 94 countries (Cystic Fibrosis Foundation). More than 60% of people with CF in the US are now adults.
Care for cystic fibrosis has changed dramatically. Medicines that target the cause of the disease, called CFTR modulators, have transformed life for many people. This guide covers seven essential facts: what causes CF, how it affects the lungs, how it is diagnosed, how it is treated, how life expectancy has changed, healthy habits and why specialist care matters.
1. Cystic Fibrosis Is a Genetic Disorder Caused by CFTR Gene Variants
CF is caused by changes (variants or mutations) in the CFTR gene. The gene carries instructions for a protein that acts like a gate for salt and water at the surface of cells. If that gate is absent or faulty, secretions lose their watery quality and turn heavy and gluey (MedlinePlus Genetics).
More than 1,700 CF-causing CFTR variants have been identified. The most common is F508del.
Inheritance and Family Risk
CF follows what geneticists call an autosomal recessive pattern: you only develop it if you receive a CF-causing variant from both your mother and your father. Someone with just one copy is a carrier and is typically healthy.
| Both Parents Are Carriers | Chance for Each Pregnancy |
|---|---|
| Child has CF | 25% |
| Child is a carrier | 50% |
| Child neither has CF nor is a carrier | 25% |
Table 1: Inheritance chances when both parents carry a CF variant (Cystic Fibrosis Foundation).
Carrier screening is offered to people planning a pregnancy or who are pregnant. Genetic counseling helps families understand their results and options.
Why Early Knowledge Matters
Finding CF early allows treatment to start before serious lung and nutrition problems develop. In the US, more than 75% of people with CF are diagnosed by age 2.
2. Cystic Fibrosis Mainly Affects the Lungs and Digestive System
How Mucus Damages the Lungs
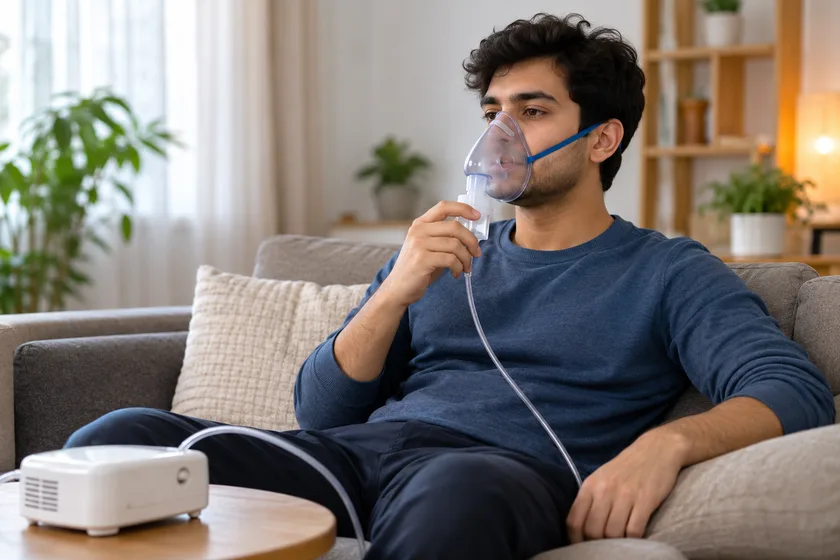
Sticky mucus blocks the small airways and gives bacteria a place to settle. The result is a cycle of infection and inflammation that can scar and widen the airways permanently (bronchiectasis). Lung disease remains the main cause of serious illness in CF (NHLBI).
Common Cystic Fibrosis Symptoms
- Persistent cough, often with thick mucus
- Wheezing or shortness of breath
- Frequent lung or sinus infections
- Nasal polyps
- Very salty-tasting skin
- Poor weight gain or growth despite a good appetite
- Greasy, bulky, foul-smelling stools
- Constipation or bowel blockage, including in newborns
- Male infertility, because the tubes that carry sperm are often missing or blocked
CF can also cause CF-related diabetes, liver disease and thin bones.
Preventing Lung Damage
Daily airway clearance, inhaled medicines, and quick treatment of infections help protect the lungs. Your CF team will tailor a plan to your needs.
3. Cystic Fibrosis Diagnosis Requires Specialized Testing
Newborn Screening
All US states screen newborns for CF with a blood test from a heel prick. A positive screen does not mean a baby has CF; it means more testing is needed.
Sweat Test
To confirm CF, doctors rely mainly on the sweat chloride test. It is painless: a little sweat is gathered from the arm and checked for chloride, and a high reading suggests CF.
| Sweat Chloride (mmol/L) | Meaning |
|---|---|
| 60 or higher | Consistent with CF |
| 30 to 59 | Intermediate; more testing needed |
| Below 30 | CF is unlikely |
Table 2: How sweat chloride results are usually interpreted.
Genetic Testing
Genetic testing identifies which CFTR variants a person has. This helps confirm the diagnosis and shows whether CFTR modulator medicines may work.
4. Treatments Can Control Symptoms, but There Is No Cure Yet
CFTR Modulator Therapies
CFTR modulators are tablets that help the faulty CFTR protein do its job more effectively. They treat the underlying cause rather than only the symptoms, but they only work for certain variants. Five modulators are currently available in the US (Cystic Fibrosis Foundation):
| Medicine | US-Approved Ages (eligibility also depends on CFTR variants) |
|---|---|
| Alyftrek (vanzacaftor/tezacaftor/deutivacaftor), once daily | 6 years and older |
| Trikafta (elexacaftor/tezacaftor/ivacaftor) | 2 years and older |
| Symdeko (tezacaftor/ivacaftor) | 6 years and older |
| Orkambi (lumacaftor/ivacaftor) | 1 year and older, with two copies of F508del |
| Kalydeco (ivacaftor) | 1 month and older |
Table 3: CFTR modulators available in the US. Eligibility depends on genetic variants.
Modulators can cause side effects, including changes in liver tests, so regular monitoring is needed.
Digestive Support and Nutrition
Because the pancreas is often blocked, most people with CF swallow enzyme capsules each time they eat so they can absorb fat and protein. A high-calorie, high-protein diet, extra salt and fat-soluble vitamins (A, D, E and K) are usually advised. A CF dietitian helps plan meals.
Daily Disease Management
- Airway clearance: chest physical therapy, vest devices, breathing techniques and exercise
- Inhaled medicines: bronchodilators, mucus thinners such as dornase alfa and hypertonic saline, and inhaled antibiotics
- Antibiotics for lung infections
- Lung transplant for some people with advanced lung disease
5. Life Expectancy Has Improved Significantly With Modern Care
Current Survival Trends
According to the Cystic Fibrosis Foundation's 2025 Patient Registry, half of people with CF born between 2021 and 2025 are predicted to live to age 66 or older, up from 65 in the 2024 report (Cystic Fibrosis Foundation). This is a population estimate, not a prediction for any one person, and it is lower for people who cannot use CFTR modulators.
Factors Affecting Life Expectancy
- Genetic variants and access to CFTR modulators
- Lung function and infections
- Nutrition and body weight
- CF-related diabetes and liver disease
- Access to specialist CF care
Living a Healthier Life With Cystic Fibrosis
Many adults with CF now study, work, have families and live full lives. Some people with CF can have children, though men often need fertility treatment.
6. Healthy Habits Support Better Cystic Fibrosis Management
Staying Physically Active
Regular exercise helps clear mucus, improves fitness and supports bone and mental health. Good choices include walking, swimming, cycling and strength training.
Preventing Infections
- Wash hands often.
- Stay up to date with vaccines, including the yearly flu shot.
- Clean nebulizers and airway equipment as instructed.
- Keep at least 6 feet (2 meters) from other people with CF, as they can pass harmful germs to each other.
- Avoid smoke and vaping.
Emotional and Social Support
Depression and anxiety are common in people with CF and their caregivers. Screening and support are part of good CF care. If breathing is a concern, our guide to mullein tea for lung health explains why herbal remedies cannot replace prescribed treatment.
7. Regular Medical Care Is Essential for Long-Term Health
Importance of Specialist Care
People with CF do best with care from an accredited CF care center, where a team of doctors, nurses, dietitians, respiratory therapists, social workers and psychologists work together. Visits are usually every 3 months for most people with CF, and infants and young children may be seen more often.
Monitoring Disease Progression
- Lung function tests (spirometry)
- Sputum or throat cultures to check for germs
- Blood tests, including diabetes and liver checks
- Growth, weight and nutrition checks
- Chest imaging and bone density scans as needed
Future Advances
Researchers are working on genetic therapies and treatments for people whose variants do not respond to current modulators.
When to See a Doctor
Contact your doctor or CF care team if you or your child has:
- A cough that is worse or brings up more mucus, or mucus that changes color
- Fever, tiredness or loss of appetite
- Weight loss or poor growth
- Greasy stools, belly pain or constipation
- Increased thirst and urination, which may suggest CF-related diabetes
- A positive newborn screen or a family history of CF
Emergency: Call 911 (or your local emergency number) for severe shortness of breath, coughing up more than a small amount of blood, chest pain with sudden breathlessness (possible collapsed lung), blue lips, confusion, or severe belly pain with vomiting and a swollen belly.
Conclusion
Cystic fibrosis is an inherited disease that results from variants in the CFTR gene. It hits the lungs and digestive system hardest and is diagnosed through newborn screening, sweat tests and genetic testing. There is no cure yet, but CFTR modulators, airway clearance, nutrition support and specialist care have helped people with CF live much longer and better lives.
Sources
- About Cystic Fibrosis (Cystic Fibrosis Foundation)
- CFTR Modulator Therapies (Cystic Fibrosis Foundation)
- 2025 Patient Registry Highlights (Cystic Fibrosis Foundation)
- Cystic Fibrosis (NHLBI)
- Cystic Fibrosis (MedlinePlus Genetics)
FAQ
1. What causes cystic fibrosis?
CF develops when a child receives a faulty CFTR gene from both parents, which leads to thick, sticky mucus.
2. Is cystic fibrosis contagious?
No. CF is a genetic condition and cannot be caught, but people with CF are kept apart from each other to avoid sharing harmful germs.
3. How is cystic fibrosis diagnosed?
Most cases are found through newborn screening, then confirmed with a sweat chloride test and genetic testing.
4. What is the life expectancy for cystic fibrosis?
According to the 2025 CF Foundation registry, half of people with CF born between 2021 and 2025 are predicted to live to 66 or older, though individual outlook varies.
5. Is there a cure for cystic fibrosis?
There is no cure yet, but CFTR modulators and other treatments greatly improve health and life expectancy for many people.
About the author
Health Researcher and Writer
Haider Nadeem is a health researcher and the writer behind Healthy Eats Guide. He writes about health problems he faces or sees people around him face, and turns careful research into practical guides. Articles are researched and drafted with AI assistance, then fact-checked and edited by Haider using cited sources such as USDA, NIH, and peer-reviewed research.
Related blogs
Mounjaro Fatigue: 5 Essential Facts to Know
Mounjaro fatigue explained: what the FDA label says, 5 key facts about tirzepatide tiredness, common causes like low intake and dehydration, and when to seek care.
9 min read
Exdensur for Asthma: 7 Essential Facts to Know
Exdensur for asthma explained: how depemokimab works, who qualifies, the twice-yearly dosing schedule, trial results, side effects and its nasal polyp status.
10 min read
Inluriyo for Breast Cancer: 6 Essential Facts to Know
Inluriyo for breast cancer: who qualifies, how imlunestrant works, the 400 mg daily dose, ESR1 testing, side effects, cost factors and warning signs.
10 min read
Newsletter
Get practical nutrition guides in your inbox
Clear food-by-food advice and diet tips for everyday eating — no spam, unsubscribe anytime.